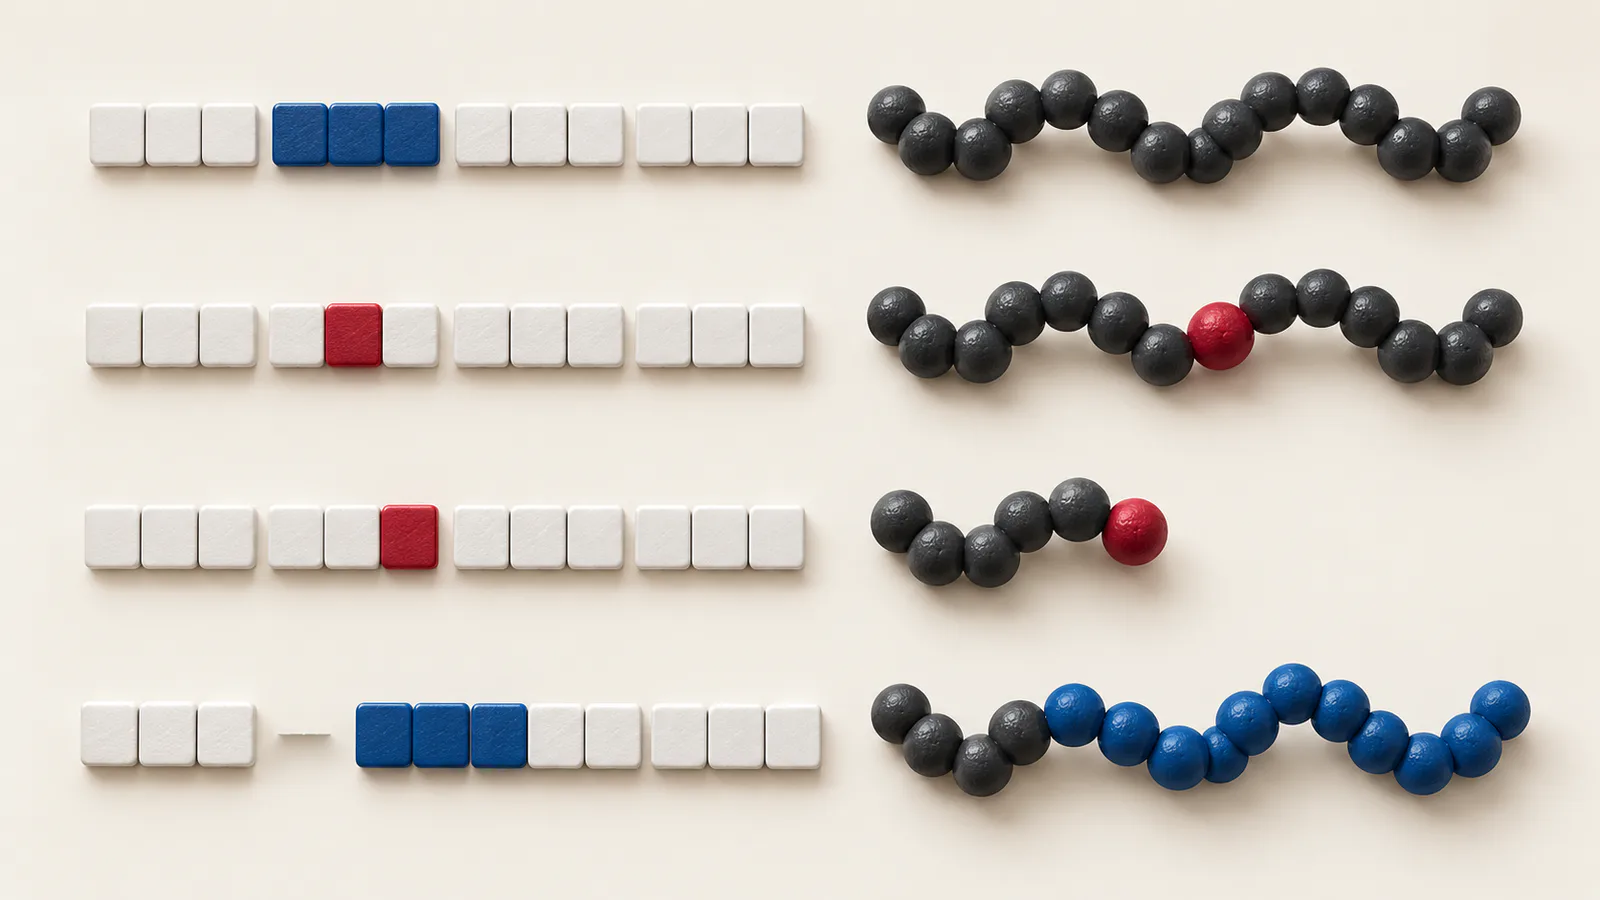
Match Each Mutation with Its Appropriate Description
Match missense with a changed amino acid, nonsense with a premature stop codon, silent with no amino-acid change, and frameshift with a shifted reading frame caused by an insertion or deletion that is not a multiple of three.
Those are the four matches most introductory questions test. The sections below show how to recognize them from DNA, RNA, or protein sequence examples.
Match each mutation with its appropriate description
Use this table for the standard four-way match.
| Mutation type | Appropriate description |
|---|---|
| Missense mutation | A nucleotide change causes a different amino acid to be incorporated |
| Nonsense mutation | A nucleotide change converts an amino-acid codon into a stop codon |
| Silent mutation | A nucleotide change does not alter the encoded amino acid |
| Frameshift mutation | An insertion or deletion shifts the reading frame downstream |
The distinction depends on the result in the coding sequence. The same physical change, such as a single-base substitution, can be silent, missense, or nonsense depending on the codon and reading frame.
What is a missense mutation?
A missense mutation changes a codon so it encodes a different amino acid.
For example, changing an mRNA codon from GAG to GUG changes glutamate to valine. The protein length may remain the same, but one residue differs.
The effect can range from negligible to severe. A conservative substitution may preserve similar chemistry, while a change at an enzyme active site or protein interface may disrupt function.
The NHGRI defines a missense mutation as a DNA change that results in a different amino acid being encoded. See the NHGRI definition.
Do not classify a substitution as missense from the DNA letters alone. Translate the original and altered codons in the correct frame first.
What is a nonsense mutation?
A nonsense mutation changes a codon for an amino acid into a stop codon.
The three stop codons in mRNA are UAA, UAG, and UGA. In a coding DNA sequence, they appear as TAA, TAG, and TGA on the coding strand.
A premature stop can produce a shortened protein. It can also trigger nonsense-mediated mRNA decay, which reduces the amount of mutant transcript before much protein is made.
The outcome depends on stop position and transcript structure. An early stop often removes more functional domains than a stop near the normal end, but molecular surveillance rules also matter.
“Nonsense” describes the coding consequence. It does not mean the variant lacks biological meaning.
What is a silent mutation?
A silent or synonymous mutation changes a codon without changing the encoded amino acid.
The genetic code is redundant. For example, GAA and GAG both encode glutamate, so a substitution between them is synonymous at the protein-sequence level.
Silent does not always mean biologically harmless. A synonymous variant can alter splicing, RNA structure, mRNA stability, codon usage, translation speed, or regulatory motifs.
A recent review summarizes evidence that synonymous variants can affect gene expression and disease mechanisms despite preserving amino-acid identity. Read the open-access review.
For a basic matching question, use “no amino-acid change.” For biological interpretation, examine effects beyond the translated sequence.
What is a frameshift mutation?
A frameshift mutation is an insertion or deletion that changes how downstream bases are grouped into codons.
Codons are read in groups of three. Adding or removing one or two bases shifts the grouping, usually changing every amino acid after the variant until a stop codon appears.
The NHGRI notes that insertions or deletions alter the reading frame unless the number of added or removed bases is a multiple of three. See the NHGRI frameshift definition.
Consider this coding sequence grouped into codons:
Original: AUG | GAA | CCU | GGC | UAAProtein: Met | Glu | Pro | Gly | StopDeleting one base changes the grouping:
Deleted: AUG | GAC | CUG | GCU | ...Protein: Met | Asp | Leu | Ala | ...The exact example is illustrative. The key clue is that all downstream codon boundaries change.
Not every insertion or deletion is a frameshift
An in-frame insertion or deletion adds or removes a multiple of three coding bases. It changes one or more amino acids but preserves the downstream reading frame.
How do you classify a mutation step by step?
Classify the physical DNA change first, then determine its consequence in the correct transcript and reading frame.
- Identify whether bases were substituted, inserted, deleted, duplicated, or rearranged.
- Confirm the reference transcript and coding strand.
- Locate the change in exon, intron, splice region, untranslated region, or intergenic sequence.
- If coding, group bases into codons from the true start of the coding sequence.
- Translate the original and altered sequence.
- Compare amino-acid identity, stop position, and downstream frame.
- Check possible splicing and transcript-level effects.
This order prevents a common mistake: assigning a protein consequence before verifying which transcript is affected.
One genomic variant can be coding in one transcript and intronic or untranslated in another.
Can a substitution cause a frameshift?
No. A simple substitution replaces one base with another and does not change sequence length, so it does not shift codon boundaries.
A substitution can be silent, missense, or nonsense within a coding region. It may also alter a splice site or regulatory sequence outside the coding region.
Complex variants can combine substitutions with insertions or deletions. In those cases, classify the complete change rather than one base in isolation.
Can an insertion or deletion avoid a frameshift?
Yes. An insertion or deletion of 3, 6, 9, or another multiple of three coding bases is in-frame.
An in-frame deletion removes amino acids while leaving later codons grouped correctly. An in-frame insertion adds amino acids without shifting the later frame.
These variants can still be damaging. Removing one critical residue from a binding pocket may matter more than changing many residues in a flexible region.
The reading-frame rule classifies the sequence consequence. It does not predict clinical importance by itself.
What other mutation descriptions should you know?
Other common descriptions include splice-site, start-loss, stop-loss, regulatory, duplication, repeat expansion, and large structural variants.
| Variant description | Meaning |
|---|---|
| Splice-site variant | Alters a sequence needed for RNA splicing and may change exon inclusion |
| Start-loss variant | Disrupts the normal translation start codon |
| Stop-loss variant | Changes the normal stop codon and may extend the protein |
| In-frame indel | Adds or removes a multiple of three coding bases |
| Regulatory variant | Alters a promoter, enhancer, or another control element |
| Duplication | Copies a sequence segment, from a few bases to a large region |
| Inversion | Reverses the orientation of a DNA segment |
| Translocation | Joins DNA segments from different genomic locations |
A splice-site change can cause exon skipping, intron retention, or use of a cryptic splice site. Our guide to identifying introns in a sequence explains how exon boundaries are established.
What is the difference between mutation type and mutation effect?
Mutation type describes the sequence change, while mutation effect describes its molecular or biological consequence.
“Single-nucleotide variant” describes a one-base change. “Missense” describes one possible coding consequence. “Loss of function” describes a broader functional outcome.
These terms should not be treated as synonyms. A missense variant may cause loss of function, gain of function, dominant-negative activity, or no measurable change.
Likewise, a frameshift often disrupts function but can occur near the end of a protein or in a transcript that is weakly expressed.
Clinical classification adds another layer, such as pathogenic, likely pathogenic, uncertain significance, likely benign, or benign. That classification requires population, functional, computational, segregation, and clinical evidence.
Practice matching examples
Apply the consequence rules to these examples before checking the answers.
| Example | Best match |
|---|---|
AAA becomes AAG, and both encode lysine | Silent mutation |
UAU becomes UAA | Nonsense mutation |
| One base is deleted from the middle of a coding region | Frameshift mutation |
GGC becomes GAC, changing glycine to aspartate | Missense mutation |
| Six coding bases are deleted | In-frame deletion, not a frameshift |
| The invariant base at an exon boundary changes | Splice-site variant |
The first four match the usual worksheet categories. The last two show why real variant annotation needs more than those four labels.
Common mistakes when matching mutation types
The most common mistake is classifying from nucleotide change alone without translating the affected codon.
Other mistakes include:
- Calling every substitution missense
- Calling every insertion or deletion a frameshift
- Treating synonymous as guaranteed benign
- Reading the template strand as if it were the coding strand
- Starting translation in the wrong frame
- Ignoring transcript isoforms
- Confusing a stop-loss variant with a nonsense variant
- Inferring disease risk from mutation class alone
Use the reference transcript, strand, exon coordinates, and coding frame whenever the problem provides them.
Key takeaways
Missense changes one amino acid. Nonsense creates a premature stop. Silent preserves the amino acid. Frameshift changes downstream codon grouping.
Substitutions can be silent, missense, or nonsense. Insertions and deletions cause frameshifts only when their coding length is not divisible by three.
Real biological interpretation also considers splicing, transcript choice, protein domain, RNA regulation, and functional evidence.
Further reading
Read another related post
STAR vs HISAT2 vs Salmon: Which Aligner Should You Use?
STAR aligns to the genome, HISAT2 uses less memory, Salmon skips alignment. What each approach means for your RNA-seq results and when each is the right call.
Research GuideWhat Is GSEA? Gene Set Enrichment Analysis Explained
Gene Set Enrichment Analysis explained: ranked gene lists, enrichment scores, NES, FDR, leading-edge genes, and an fgsea tutorial in R.
Research GuideRNA-Seq Sample Preparation: RNA Extraction to FASTQ Files
RNA-seq sample preparation from RNA extraction and quality checks through library preparation, Illumina sequencing, base calling, and FASTQ files.